Device entries stall on a wrong affirmation code, a firm on an import alert, or a tariff line nobody rechecked. We file them as
one licensed broker handling FDA data, duty exposure, and device freight for OEMs and distributors.
Medical device logistics fails at the data layer. Every device line carries an FDA product code and affirmation of compliance codes in ACE, and a device on the wrong import alert or with a stale listing number is detained no matter how clean the CBP entry looks.
A licensed customs broker on our team files the CBP entry and the FDA PGA message set together. Where your company also holds FDA initial importer duties, that program and the entry data run from the same records.
Duty is the second exposure. Most chapter 90 lines are free at MFN, so the cost sits in chapter 99: Section 301 on China, the forced-labor tariffs in force since July 24, 2026, and an open Section 232 investigation on medical products.
$13,785
FY2027 FDA registration fee
100%
Section 301 on China syringes and needles
12.5%
Top forced-labor 301 rate, no device exemption
Free Device Entry Review
Talk to a Licensed Broker
Quick 30-min review of your device codes, import alerts, and tariff lines. No obligation.
We reply within 1 business day · Your data stays private.
Capabilities
Our Medical Device Customs Brokerage Capabilities
Six operations that keep medical device freight admissible, correctly dutied, and handled the way the manufacturer's labeling requires.
01
FDA Entry Data for Devices
Product codes and affirmation of compliance codes mapped per SKU before the first entry.
DEV, DFE and LST registration codes
PM# premarket numbers from the listing
DI (UDI) and KIT where they apply
Form FDA 2877 codes for radiation products
02
Import Alert Screening
Suppliers and product codes checked against the device import alerts before booking.
Why CargoTrans for Medical Device Customs Brokerage?
The FDA message set and the CBP entry are filed by the same licensed broker, so the affirmation codes, product code, and HTS line on each device agree before FDA screens them.
Suppliers are checked against device import alerts such as 99-34, 89-08, and 99-49 before booking, when switching a source still costs less than a detention.
Chapter 99 exposure is modeled per HTS line and origin, including Section 301 on China and the forced-labor tariffs, so a landed cost does not change after the goods arrive.
Lithium battery classification, labeled storage conditions, and lot and serial records are set at booking, which is where OEM quality agreements expect them to live.
Our Process
How Our Medical Device Customs Brokerage Process Works
01
Catalog and Code Mapping
Map product codes, AofC codes, and HTS per device.
02
Supplier and Alert Check
Screen manufacturers and codes against device import alerts.
03
Duty and Lane Setup
Model chapter 99 duty and book compliant device lanes.
04
File and Clear
Transmit CBP entry and FDA data as one filing.
05
Store, Trace, Report
Hold per labeling and keep lot records audit-ready.
A device shipment can clear CBP and still sit on an FDA hold over one mismatched listing number or a supplier on an import alert. We close that gap before the freight leaves origin.
CargoTrans files device entries, prices the duty, and moves the freight as one job.
Free 30-minute device entry and tariff review
Affirmation of compliance codes mapped per device
Section 301 and forced-labor exposure priced per HTS line
Lithium documentation and labeled storage conditions set at booking
A CargoTrans licensed broker will review your device catalog, suppliers, and tariff lines and flag any FDA or duty exposure.
Protected by reCAPTCHA. We respond within 1 business day. No spam, ever.
Control Tower
Medical Device Entries, Live in the Control Tower
Your broker sees each device entry's FDA disposition as it posts and flags it in the Captain Control Tower beside the CBP release, so a May Proceed, a document request, or a Notice of FDA Action reaches you without waiting on a broker email days later.
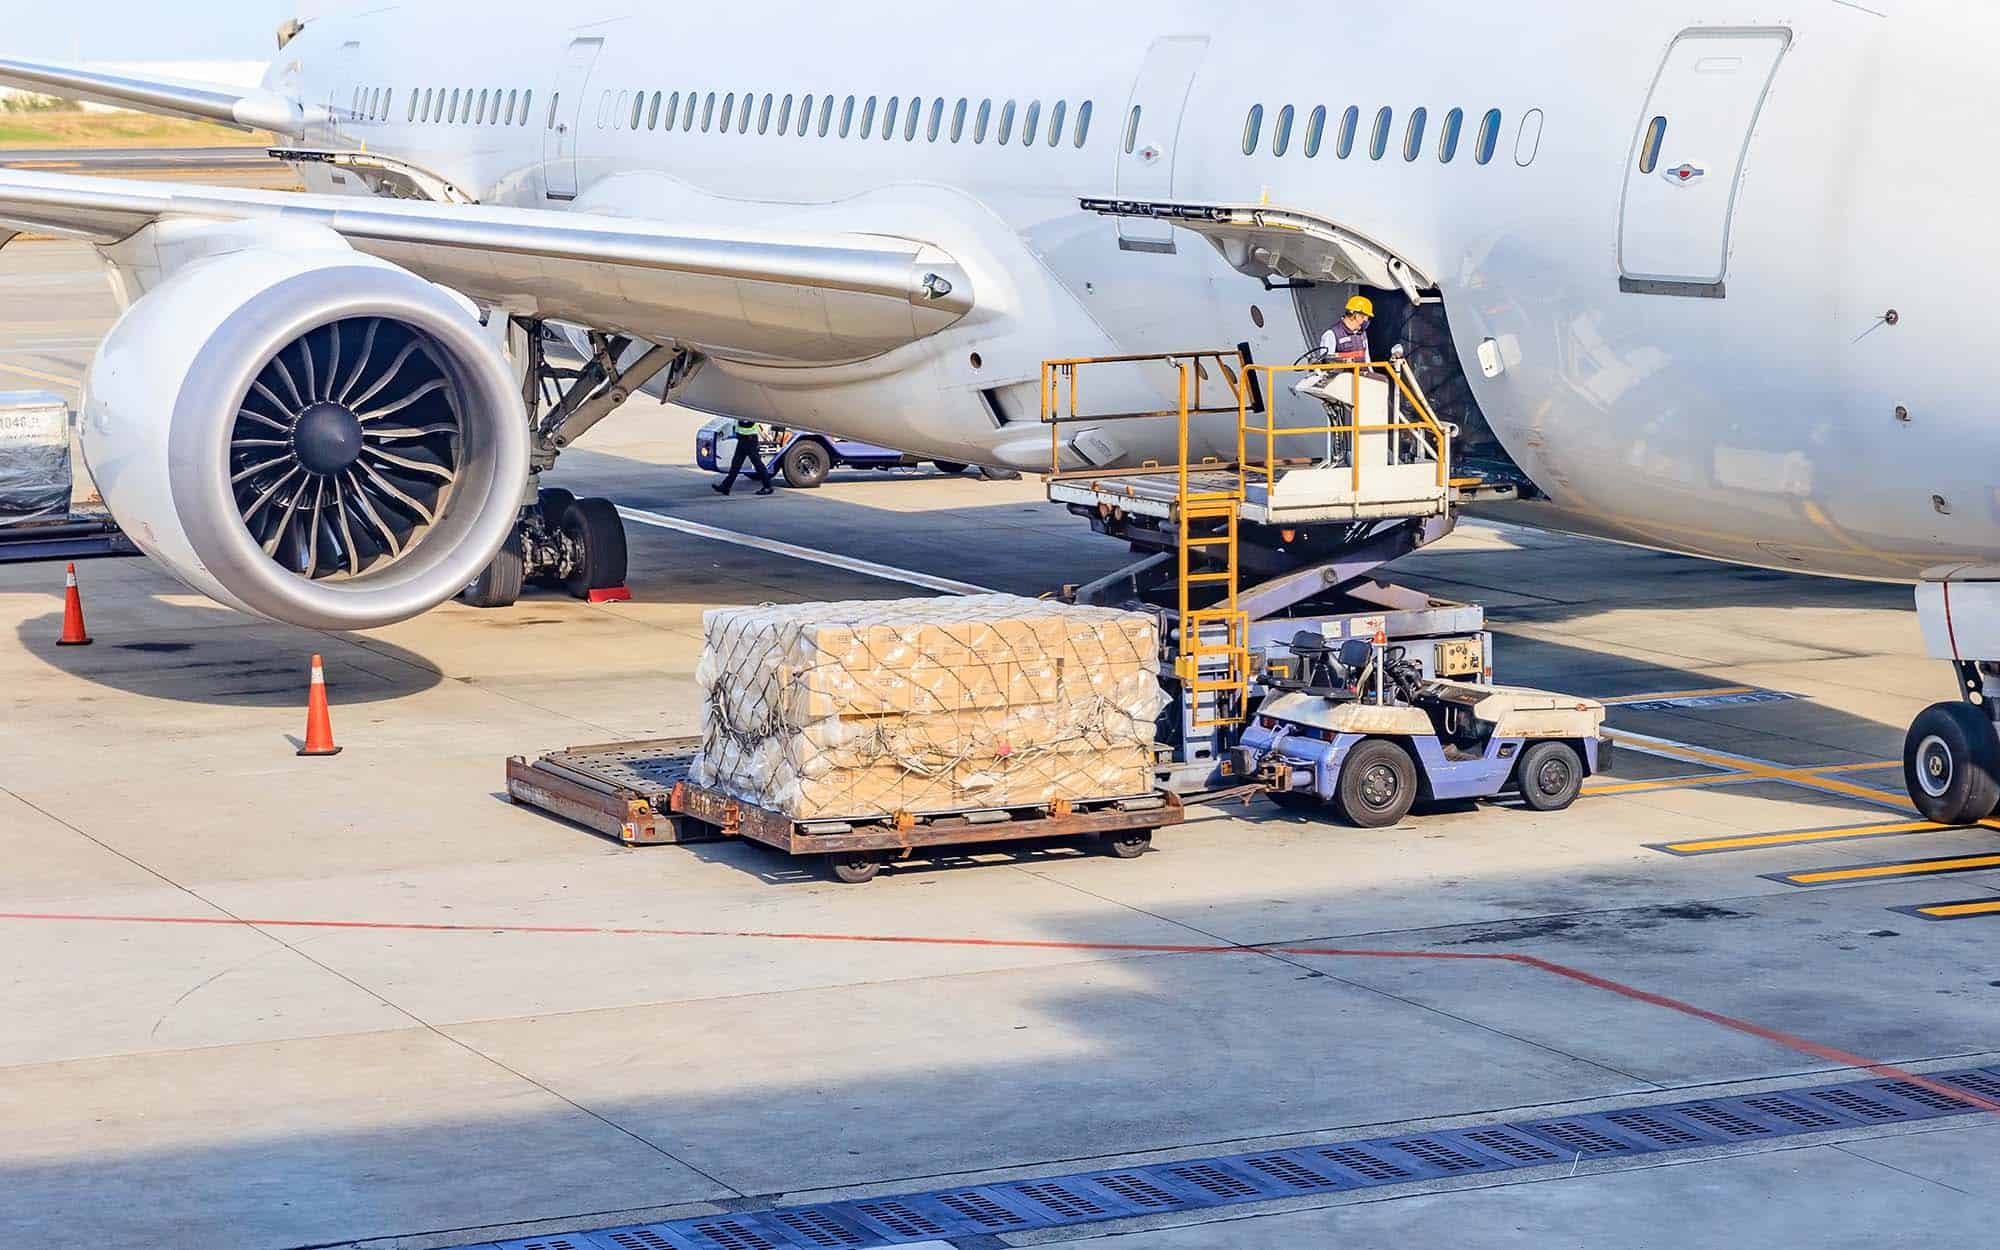
Alerts flag held lines, battery exceptions, and temperature-sensitive loads early, so a device shipment waiting on one missing premarket number is fixed while the aircraft is still in the air.
Every affirmation code, product code, HTS line, and FDA disposition is stored with the entry, so a CBP audit, an FDA document request, or a Section 301 refund claim is answered from one file rather than rebuilt.
Lot and serial data carried from your records into the entry file stay with the entry record, which gives your quality team the traceability that MDR reports and corrections and removals under 21 CFR 806 depend on.
What does a customs broker for medical devices handle that a general broker does not?
A general broker files the CBP entry and pays the duty. A device entry adds a second filing that decides whether the goods can be sold at all: the FDA PGA message set in ACE, with an FDA product code and affirmation of compliance codes on every line. The broker has to know which codes apply to which device, which suppliers sit on an FDA import alert, and which premarket number appears on the listing record, because FDA screening compares them. Duty is the other difference. Most chapter 90 lines are free at MFN, so the real cost comes from chapter 99 measures such as Section 301 on China and the forced-labor tariffs, which change by origin and HTS line. Add lithium battery rules and labeled storage conditions, and a device entry needs a broker who treats FDA data, tariff exposure, and freight handling as one job.
Which FDA affirmation of compliance codes go on a medical device entry?
FDA's current ACE guide, effective March 10, 2026, lists the device codes. DEV carries the foreign manufacturer's registration number, DFE the foreign exporter's, and DDM a US manufacturer's. LST is the device listing number, PM# the premarket number (510(k), PMA, De Novo, HDE and similar), and FDA says it should match the number on the listing record. IDE covers investigational devices, DI carries the UDI device identifier, KIT flags imported kits of finished devices, and LWC covers electrode lead wires and patient cables. Radiation-emitting products add the Form FDA 2877 declarations as RA, RB, RC and RD codes. There is no code for the initial importer's own registration, and REG is a drug establishment registration code, not a device code. FDA states that declaring a code it does not list results in an entry rejection, so the mapping is done per SKU before the first shipment.
How does UDI data affect medical device customs clearance?
The unique device identifier now matters at the border, not only on the label. FDA's ACE guide includes a DI affirmation of compliance code that carries the device identifier portion of the UDI, and Import Alert 99-49, published June 22, 2026, allows detention without physical examination of devices that lack a UDI or a matching GUDID record. UDI labeling and GUDID submission are the labeler's duties under 21 CFR 801 and 830, not a customs task, but a gap there detains the importer's shipment all the same. FDA's quick reference guide lists DI as optional on a standard device entry (DEV, DFE and LST are mandatory), but transmitting it lets FDA match the shipment to GUDID, which is what Import Alert 99-49 screens. The same identifier appears downstream: MDR reports and corrections and removals reports both carry the UDI. We collect device identifiers with the catalog, check them against GUDID before the first entry, and flag any SKU whose label and database record disagree.
Which FDA import alerts affect medical device importers?
Import alerts let FDA detain goods without physical examination, and several target devices directly. Import Alert 99-34 covers drugs and devices from firms without a valid registration or listing. 89-08 covers devices without an approved PMA or IDE, or without 510(k) clearance. 89-04 targets firms that have not met device quality system requirements, and 89-16 firms that refused an FDA foreign inspection. 99-49, published June 22, 2026, covers devices without a unique device identifier or a matching GUDID record. Product-specific alerts include 80-04 for surgeon's and patient examination gloves, 89-18 for filtering facepiece respirators, 79-01 for bandages and gauze with microbiological contamination, and 76-01 for medical instruments from Pakistan. A supplier or product code on one of these lists turns every entry into a detention, so we check the lists before the purchase order ships, not after the goods land.
How much is the FDA device establishment registration fee and who pays it?
Every registered device establishment pays an annual fee, with no small-business discount on registration. For FY2026 (October 1, 2025 to September 30, 2026) it was $11,423. For FY2027, which starts October 1, 2026, FDA set it at $13,785 in a Federal Register notice published July 30, 2026. Registration is renewed each year between October 1 and December 31. Foreign manufacturers, foreign exporters, specification developers, and initial importers all register and pay. Foreign manufacturers and specification developers also list their devices; an initial importer meets its listing obligation by identifying the manufacturer. Registration and listing matter at the border because 21 CFR 807.40(c) bars importing a device unless it is listed and made at a registered foreign establishment, which is also what Import Alert 99-34 enforces. We confirm the status of every manufacturer in your supply base before the first entry.
What Section 301 duties apply to medical devices from China?
USTR's four-year review, finalized September 18, 2024, raised several medical lines. Syringes under HTS 9018.31.00 and tubular metal and suture needles under 9018.32.00 went to 100% in 2024. Medical and surgical rubber gloves under 4015.12.10 went to 50% in 2025 and 100% from January 1, 2026. Surgical and non-surgical N95 respirators, other textile respirators, and textile face masks under the 6307.90.98 subheadings reached 50% in 2026. Those rates stack on top of MFN duty and on top of the separate forced-labor tariff that took effect July 24, 2026. A set of 178 legacy exclusions runs to November 10, 2026, and USTR opened a second four-year review of the China actions on May 6, 2026, so the rates can move again. We price each line by HTS and origin, and whether an exclusion covers a given device is checked against the annex, never assumed.
Do the Section 301 forced-labor tariffs exempt medical devices?
No. USTR's forced-labor action covers 60 economies from 12:01 a.m. ET on July 24, 2026, at 10% or 12.5% depending on the trading partner. For major device sources: the EU is 10% net of MFN, Mexico, Canada, the UK and Malaysia 10%, and China, Singapore, Israel, Costa Rica and the Dominican Republic 12.5%, while Japan, Switzerland and South Korea are 12.5% net of MFN. Commenters asked for a medical device exemption and USTR did not add one, and it declined to create an exclusion process. Three carve-outs matter: USMCA duty-free goods of Canada and Mexico are exempt, articles subject to Section 232 tariffs are exempt, and goods properly entered under chapter 98 are exempt, except 9802.00.40, .50, .60 and 9802.00.80, where the tariff applies only to the value added abroad. For a device OEM, that makes USMCA qualification and chapter 98 eligibility worth checking line by line, since the chapter 90 MFN rate is often zero.
Is there a Section 232 tariff on medical devices?
Not yet. Commerce opened a Section 232 investigation into personal protective equipment, medical consumables, and medical equipment, including devices, on September 2, 2025. The notice named masks, N95s, gloves, gowns, syringes, needles, infusion pumps, IV bags, catheters, bandages, diagnostic reagents, wheelchairs, crutches and hospital beds. Comments closed October 17, 2025. The statute gives Commerce 270 days to report, which put the report on or about May 30, 2026, and the President 90 days after that to act. As of September 2026 no proclamation has been issued on medical products, while 232 actions have been taken on semiconductors, pharmaceuticals, critical minerals and other sectors. Whether Commerce has delivered its report is not public. If a proclamation issues, goods it covers would drop out of the forced-labor 301 tariff, so we track the docket and our Section 232 tariff consulting team models both scenarios.
When can a medical device enter duty-free under HTS 9817.00.96?
HTS 9817.00.96 implements the Nairobi Protocol and is free of duty. It covers articles specially designed or adapted for the use or benefit of the blind or other physically or mentally handicapped persons, and their parts, where the person has a permanent or chronic impairment that substantially limits a major life activity. The exclusions are what trip importers up: articles for acute or transient disability, spectacles, dentures and cosmetic articles for people who are not substantially disabled, therapeutic and diagnostic articles, and medicine are all outside it. Parts of braces and artificial limb prosthetics are excluded from the parts clause. The claim matters more than it used to, because goods properly entered under chapter 98 are exempt from the forced-labor 301 tariff. We review design documents and intended use before claiming it, because a claim that CBP later rejects reopens the full chapter 99 duty.
How do you ship medical devices that contain lithium batteries?
A device with its battery installed ships as lithium ion batteries contained in equipment, UN3481, or lithium metal batteries contained in equipment, UN3091, under 49 CFR 173.185. Small consignments of batteries in equipment can qualify for relief from the lithium battery mark, and the rule has specific provisions for medical devices: up to two replacement cells or batteries for a medical device may go as cargo on a passenger aircraft with PHMSA approval when the destination has no daily cargo service, and medical-device lithium ion cells can be excepted from the state-of-charge limit under stated conditions. Airline acceptance adds its own layer, so we confirm each carrier's rules before booking. Urgent device replenishment usually moves on air freight, which is exactly where battery documentation errors cause refused tenders, so classification is fixed at booking rather than at the terminal.
How do you handle temperature-sensitive and sterile medical devices?
FDA does not publish a transport temperature rule for devices. The requirement comes from the manufacturer's labeling and from its quality system. The Quality Management System Regulation, in effect since February 2, 2026 and incorporating ISO 13485:2016, requires manufacturers to control labeling and packaging integrity during storage, handling and distribution, and ISO 13485 clause 7.5.11 covers product preservation. OEMs pass those duties to their logistics providers through quality agreements and supplier audits, and a sterile barrier is part of that packaging integrity. We book carriers to the labeled range, protect sterile packaging from crush and moisture, and arrange warehouse and distribution storage matched to the manufacturer's labeled conditions, per shipment. If your quality agreement sets temperature logging, excursion reporting, or audit rights, those terms are built into the lane before the first shipment moves.
What happens when FDA detains a medical device shipment?
FDA issues a Notice of FDA Action marked Detained, which works as the notice of detention and hearing. The response is due within 10 business days, and the notice usually shows a date 20 calendar days out; FDA can extend it on a justified request. The response is evidence: a registration or listing record, a 510(k) or PMA number, corrected affirmation codes, or labeling. If the problem can be fixed, a reconditioning proposal goes in on Form FDA 766. With no response, FDA can refuse admission, and refused goods must be exported or destroyed under CBP supervision within 90 days. Because we file the FDA data and hold the supplier records, we see the notice when it posts, tell you the cause, and prepare the response from documents already in the file, which is what keeps a detention from becoming a refusal and a re-export bill.
How do the importer, specification developer, and US Agent roles differ?
Under 21 CFR 807.3(g), the initial importer furthers the marketing of a foreign manufacturer's device to the party who makes final delivery or sale, without repackaging or changing its labeling. Relabel or repack, and the rules treat you as a manufacturer. A US brand owner that sets specifications for a device a contract manufacturer builds abroad is a specification developer: it registers, lists, pays the fee, and falls under the QMSR. The US Agent, required under 807.40(b) for each foreign establishment, is the foreign firm's FDA contact and does not take on importer duties. Foreign manufacturers must also identify each importer and each person who offers the device for import, brokers included, in their registration. For device tracking under 21 CFR 821, brokers and warehousers are expressly not the importer. We map which role each entity in your chain holds before the first entry.
What do you need to quote medical device logistics and brokerage?
Start with the device catalog: FDA product codes, device class, the 510(k), PMA or De Novo number on each listing, the registration numbers of each foreign manufacturer and exporter, and UDI device identifiers where you have them. Then the trade data: HTS classification, country of origin, and any USMCA or chapter 98 claims you make today. For freight, we need lanes, monthly volumes, which devices contain lithium batteries and in what configuration, labeled storage temperatures, sterile items, and any OEM quality agreement terms. With that we map affirmation codes per SKU, screen suppliers against the device import alerts, model Section 301 and forced-labor exposure per line, and quote brokerage, freight and storage separately so the fixed and per-entry costs are visible. Gaps in the catalog are normal; finding them before the first entry is part of the review.